Keyue is based in Hangzhou, China, with the correspondent for the submission listed as King Laser Medtech Co., Ltd., based in Changchun. The applicant contact was Zhao Li; the correspondent contact was Jinjing Sui. The company did not pursue a predetermined change control plan authorization, meaning any future modifications to the device will require separate regulatory submissions rather than operating under a pre-approved framework for design changes.
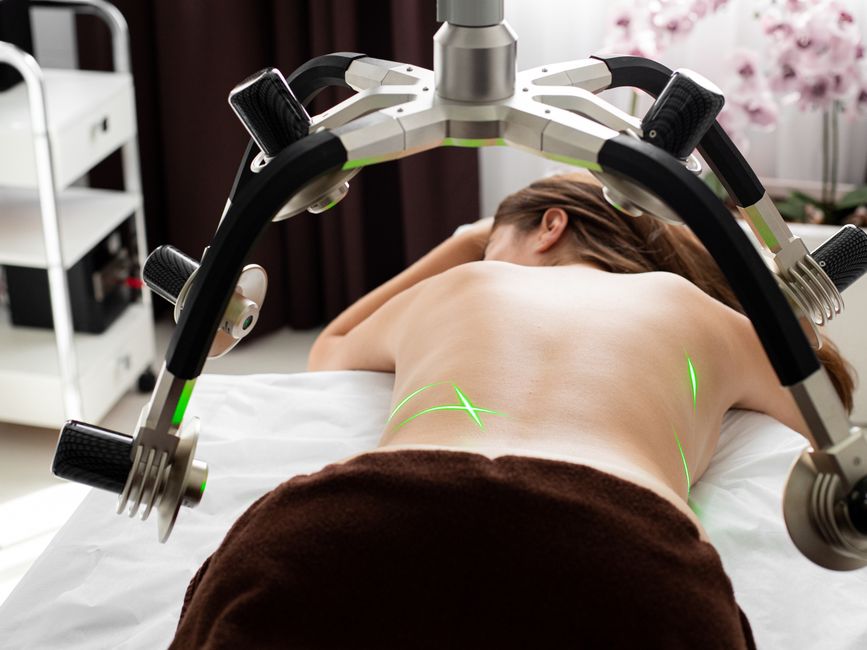
The dual-wavelength approach matters for practices because Nd:YAG and alexandrite lasers address different clinical needs. Nd:YAG penetrates deeper into tissue and works on darker skin tones, while alexandrite operates at a shorter wavelength suited to lighter skin and superficial targets. Combining both in one platform reduces the capital laser footprint compared to buying two separate systems, though the document does not specify pricing, exact parameters, or intended clinical indications beyond the general surgical classification.
The 510(k) pathway means the FDA did not conduct a full safety and efficacy review; instead, it confirmed the LPDuo performs substantially as its predicate device does. The document does not identify which specific predicate device Keyue cited, nor does it detail the clinical data or bench testing submitted to support the equivalence claim. That information typically appears in the full 510(k) summary, which is not included in this FDA database entry.
The FDA handled the review directly rather than using a third party. The device is not classified as a combination product, so it does not combine drug and device elements or other regulated components that would trigger additional oversight pathways.
For practices evaluating the LPDuo, the substantial equivalence determination signals that the FDA found no new safety or performance concerns compared to existing dual-wavelength or single-wavelength laser platforms already on the market. The traditional 510(k) pathway is faster than De Novo or PMA routes, but it also means less granular FDA scrutiny of the specific clinical claims or performance data. Practices should request the full 510(k) summary from Keyue or King Laser Medtech to understand which predicate device was used and what testing supported the equivalence claim.
Without a predetermined change control plan, Keyue cannot modify the device design, materials, or performance parameters without filing a new 510(k) or other regulatory submission. This protects the cleared status but also constrains the company's ability to iterate quickly based on field feedback or manufacturing improvements.
The decision date of October 13, 2026, makes this a recent clearance. Practices considering whether to buy the device should verify that it is actually available for purchase and that Keyue or King Laser Medtech has established distribution and support channels in the U.S. market. The FDA database entry does not confirm commercial availability or launch timeline.
Business information, not medical or legal advice. Verify regulatory status at FDA.gov.